Integrated Rate Law Calculator
Integrated rate laws for zero, first and second order: find [A] at time t, k, t or the half-life, or paste readings to see which plot is straight.
Calculator
- [A] at time t
- 0.3135 M
- Half-life The same from any starting concentration, which is the signature of first order.
- 5.975 h
- Left at time t
- 31.3 %
- Rate at time t −d[A]/dt = k[A]ⁿ at the moment t, which falls as A is used up except at zero order.
- 0.03636 M h⁻¹
- [A]
- ln[A] against t
Working
- First order: ln[A] = ln[A]₀ − kt, so [A] = [A]₀e^(−kt), and t½ = ln 2/k.
- kt = 0.116 x 10 = 1.16
- [A] = 1 x e^(−1.16) = 0.31349 M
- t½ = ln 2 / 0.116 = 5.9754 h
- A first order half-life does not depend on [A]₀.
Here k is defined by −d[A]/dt = k[A]ⁿ. A rate written as −(1/a) d[A]/dt for aA → products has a k that is 1/a of this one.
Citing this tool
Last updated . Add the date you accessed it as well, which a citation of a page that can change asks for. If a specific result matters, cite the permalink from the tool’s share row instead of this page: it reproduces the exact parameters.
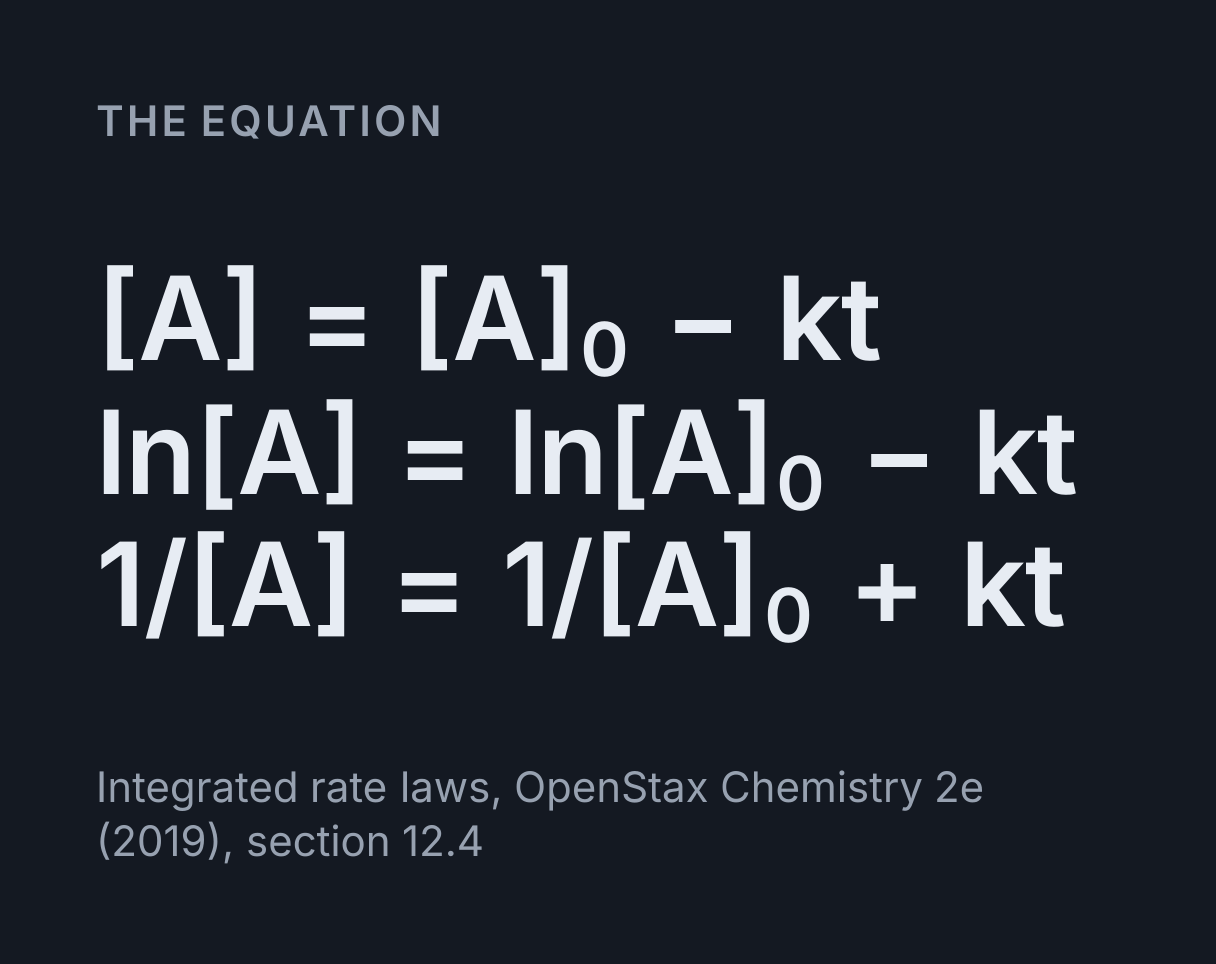
The equation
Integrated rate laws, OpenStax Chemistry 2e (2019), section 12.4
The integrated rate laws for zero, first and second order
An integrated rate law gives the concentration of a reactant A at time t from its starting
concentration [A]₀ and the rate constant k. For zero order it is [A] = [A]₀ − kt,
for first order ln[A] = ln[A]₀ − kt, which is [A] = [A]₀e^(−kt), and
for second order 1/[A] = 1/[A]₀ + kt. Each comes from integrating the rate law
−d[A]/dt = k[A]ⁿ with n = 0, 1 or 2, and each has its own half-life and its own
straight-line plot.
| Order | Rate law | Integrated form | Half-life | Straight plot | Units of k |
|---|---|---|---|---|---|
| Zero | rate = k | [A] = [A]₀ − kt | t½ = [A]₀/(2k) | [A] against t, slope −k | M s⁻¹ |
| First | rate = k[A] | ln[A] = ln[A]₀ − kt | t½ = ln 2/k | ln[A] against t, slope −k | s⁻¹ |
| Second | rate = k[A]² | 1/[A] = 1/[A]₀ + kt | t½ = 1/(k[A]₀) | 1/[A] against t, slope +k | M⁻¹ s⁻¹ |
How to use the calculator
Solve the rate law is for a question that gives you the order. Pick it, choose what to solve for, and fill in the other three: [A] after a time, the time to reach a concentration, the rate constant from two concentrations and a time, the starting concentration, or k from a half-life. The half-life comes with every answer, and two plots show the decay and the same decay on the order’s straight-line axis. Pick one concentration unit and one time unit; k is printed in the unit its order gives it.
Find the order from data is for a table of readings. Paste times and concentrations, one pair per line, and the calculator fits a least-squares straight line to each of [A], ln[A] and 1/[A] against t. The plot with the highest R² names the order, its slope gives k, and its intercept gives [A]₀. A button carries the fitted order, k and [A]₀ back into the solver, so a follow-up question about the same reaction takes one tap.
Worked example: a first order decay
Hydrogen peroxide decomposing in OpenStax’s Example 12.7 is first order with k = 0.116 h⁻¹, starting from 1.000 M, after 10 h: kt = 0.116 × 10 = 1.16, so [A] = 1.000 × e^(−1.16) = 0.3135 M, and 31 percent of the peroxide is left. The half-life does not depend on the starting amount: t½ = ln 2/0.116 = 5.975 h. These are the calculator’s defaults, so the numbers above are the ones it opens with.
Worked example: the order of a reaction from data
The data mode opens with OpenStax’s Example 12.9, the dimerisation of butadiene, with concentrations falling from 1.00 × 10⁻² M to 2.08 × 10⁻³ M over 6200 s. Fitting all three plots gives R² = 0.8238 for [A], 0.9506 for ln[A] and 1.0000 for 1/[A], so only the second order plot is straight. Its slope is the rate constant, k = 0.06144 M⁻¹ s⁻¹, which the book rounds to 0.0614 M⁻¹ s⁻¹, and at the fitted starting concentration the half-life is 1629 s.
What the half-life says about the order
Only a first order reaction has a constant half-life, which is why radioactive decay is
described by one; the half-life calculator works
through that case in half-lives. At zero order t½ = [A]₀/(2k), so each half-life is
half as long as the one before. At second order t½ = 1/(k[A]₀), so each is twice
as long. Watching successive half-lives in a table of readings is a quick check of the order
before any fitting.
The rate constant itself changes with temperature, and the Arrhenius equation calculator finds how much from the activation energy. The collision theory simulator shows why so few collisions react. For a straight-line fit with its uncertainties, residuals and a line through the origin, use the linear regression calculator.
What this does not cover
- The stoichiometric factor. Here k is defined by
−d[A]/dt = k[A]ⁿ. IUPAC defines the rate ofaA → productsas−(1/a) d[A]/dt, and with that definition k is 1/a of the value found here. For 2A → P a question may mean either, so check which one your course uses. - Two reactants. A rate law of
k[A][B]with unequal starting concentrations integrates to a different form. With B in large excess it becomes pseudo-first order in A, and the k found here is then k[B]. - Reversible, consecutive and fractional order reactions. Only the three simple orders of a single reactant going to completion are modelled.
- How good the fit is. R² ranks the three plots; it is not an uncertainty on k, and fitting ln[A] or 1/[A] weights the readings unevenly, giving the late, low concentrations more or less say than the early ones.
Common mistakes
- Using the first order half-life for every order.
t½ = ln 2/kholds only at first order. - Getting the sign of the slope wrong. [A] and ln[A] fall, so k is minus their slope; 1/[A] rises, so k is its slope.
- Mixing units. t and k must use the same time unit, and at zero and second order the same concentration unit as [A].
- Deciding the order from a short run. Over less than half the reaction all three plots are close to straight, so the calculator warns when the readings stop that early.
- Letting a zero order concentration go negative.
[A] = [A]₀ − ktstops at zero, when A runs out att = [A]₀/k.
Common questions
What is an integrated rate law?
It is the rate law integrated over time, giving the concentration of a reactant at any time t from its starting concentration and the rate constant. For zero order [A] = [A]₀ − kt, for first order ln[A] = ln[A]₀ − kt, and for second order 1/[A] = 1/[A]₀ + kt. The differential rate law −d[A]/dt = k[A]ⁿ says how fast A is used at one moment; the integrated form says how much is left after a given time.
How do you find the order of a reaction from concentration and time data?
Plot [A], ln[A] and 1/[A] against time and see which one is a straight line: [A] for zero order, ln[A] for first order, 1/[A] for second order. The slope of the straight one gives k, minus the slope for the first two and the slope itself for 1/[A]. The readings need to cover well over half the reaction, because over a short run all three plots look nearly straight.
How do you calculate the rate constant from a half-life?
At first order k = ln 2/t½, with no concentration needed: a half-life of 2.16 × 10⁴ s gives k = 0.693/21600 = 3.21 × 10⁻⁵ s⁻¹. At zero order k = [A]₀/(2t½), and at second order k = 1/([A]₀t½), so those two also need the starting concentration the half-life was measured from.
What is the half-life of a second order reaction?
It is t½ = 1/(k[A]₀), so it depends on the starting concentration and doubles with each half-life that passes. For butadiene dimerising with k = 0.0576 L mol⁻¹ min⁻¹ from 0.200 M, t½ = 1/(0.0576 × 0.200) = 86.8 min, and the next half-life, from 0.100 M, is 174 min.
What are the units of the rate constant for each order?
They are concentration to the power 1 − n per unit time, so M s⁻¹ (mol L⁻¹ s⁻¹) at zero order, s⁻¹ at first order and M⁻¹ s⁻¹ (L mol⁻¹ s⁻¹) at second order. The units of a rate constant therefore tell you the order, and a first order k is the only one that does not depend on the concentration unit.
Why does a zero order reaction stop?
Because the reactant runs out. A zero order rate does not depend on [A], so [A] = [A]₀ − kt falls in a straight line and reaches zero at t = [A]₀/k, where the law stops applying. Zero order is usually seen when something other than A limits the rate, such as a saturated catalyst surface, and it turns into another order once [A] is low enough.